
Nur für die Forschung bestimmt
EED226 DNA/RNA Synthesis Inhibitor
Kat.-Nr.: S8496
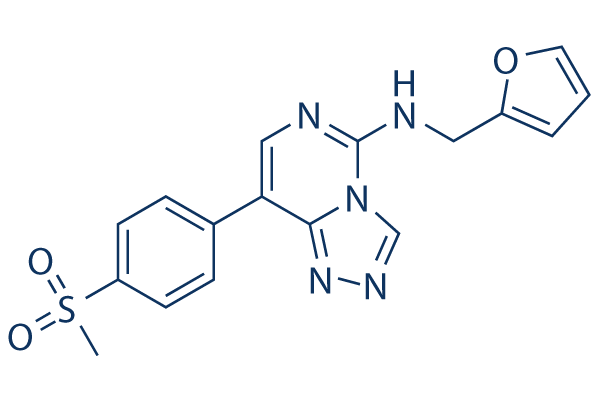
Chemische Struktur
Molekulargewicht: 369.40
Qualitätskontrolle (Quality Control)
- In Nature Medicine für seine erstklassige Qualität zitiert
- COA
- SDS
- Datenblatt
| Verwandte Targets | HDAC JAK BET Histone Methyltransferase PKC PARP HIF PRMT EZH2 AMPK |
|---|---|
| Andere DNA/RNA Synthesis Inhibitors | CX-5461 (Pidnarulex) SCR7 Favipiravir (T-705) RK-33 BMH-21 Carmofur Triapine (3-AP) YK-4-279 Halofuginone Tegafur (FT-207) |
Zellkultur, Behandlung & Arbeitskonzentration
(Cell Culture, Treatment & Working Concentration)
| Zelllinien | Assay-Typ | Konzentration | Inkubationszeit | Formulierung | Aktivitätsbeschreibung | PMID |
|---|---|---|---|---|---|---|
| KARPAS422 | Antiproliferative assay | up to 14 days | Antiproliferative activity against human KARPAS422 cells harboring monoallelic Y641N EZH2 mutation assessed as reduction in cell viability measured every 3 to 4 days up to 14 days by Beckman Coulter-based method, IC50 = 0.08 μM. | 28092155 | ||
| G401 | Function assay | 48 hrs | Inhibition of EED in human G401 cells assessed as reduction in global H3K27me3 level after 48 hrs by ELISA, IC50 = 0.22 μM. | 28092155 | ||
| KARPAS422 | Antitumor assay | 300 mg/kg | 34 days | Antitumor activity against human KARPAS422 cells xenografted in Balb/C nude mouse assessed as tumor regression at 300 mg/kg, po BID for 34 days | 28092155 | |
| KARPAS422 | Antitumor assay | 1.5 to 40 mg/kg | 2 weeks | Antitumor activity against human KARPAS422 cells xenografted in Balb/C nude mouse assessed as reduction in tumor volume at 1.5 to 40 mg/kg, po BID for 2 weeks | 28092155 | |
| Klicken Sie hier, um weitere experimentelle Zellliniendaten anzuzeigen | ||||||
Chemische Informationen, Lagerung & Stabilität (Chemical Information, Storage & Stability)
| Molekulargewicht | 369.40 | Formel | C17H15N5O3S |
Lagerung (Ab Erhaltdatum) | |
|---|---|---|---|---|---|
| CAS-Nr. | 2083627-02-3 | SDF herunterladen | Lagerung von Stammlösungen |
|
|
| Synonyme | N/A | Smiles | CS(=O)(=O)C1=CC=C(C=C1)C2=CN=C(N3C2=NN=C3)NCC4=CC=CO4 | ||
Löslichkeit (Solubility)
|
In vitro |
DMSO
: 73 mg/mL
(197.61 mM)
Water : Insoluble Ethanol : Insoluble |
Molaritätsrechner
|
In vivo |
|||||
In-vivo-Formulierungsrechner (Klare Lösung)
Schritt 1: Geben Sie unten die Informationen ein (Empfohlen: Ein zusätzliches Tier einplanen, um Verluste während des Experiments auszugleichen)
Schritt 2: Geben Sie die In-vivo-Formulierung ein (Dies ist nur der Rechner, nicht die Formulierung. Bitte kontaktieren Sie uns zuerst, wenn im Abschnitt Löslichkeit keine In-vivo-Formulierung angegeben ist.)
Berechnungsergebnisse:
Arbeitskonzentration: mg/ml;
Methode zur Herstellung der DMSO-Stammflüssigkeit: mg Wirkstoff voraufgelöst in μL DMSO ( Konzentration der Stammflüssigkeit mg/mL, Bitte kontaktieren Sie uns zuerst, wenn die Konzentration die DMSO-Löslichkeit der jeweiligen Wirkstoffcharge überschreitet. )
Methode zur Herstellung der In-vivo-Formulierung: Nehmen Sie μL DMSO Stammflüssigkeit, fügen Sie als Nächstes hinzuμL PEG300, mischen und aufklären, fügen Sie als Nächstes hinzuμL Tween 80, mischen und aufklären, fügen Sie als Nächstes hinzu μL ddH2O, mischen und aufklären.
Methode zur Herstellung der In-vivo-Formulierung: Nehmen Sie μL DMSO Stammflüssigkeit, fügen Sie als Nächstes hinzu μL Maisöl, mischen und aufklären.
Hinweis: 1. Bitte stellen Sie sicher, dass die Flüssigkeit klar ist, bevor Sie das nächste Lösungsmittel hinzufügen.
2. Achten Sie darauf, die Lösungsmittel in der richtigen Reihenfolge hinzuzufügen. Sie müssen sicherstellen, dass die im vorherigen Schritt erhaltene Lösung klar ist, bevor Sie das nächste Lösungsmittel hinzufügen. Physikalische Methoden wie Vortexen, Ultraschall oder ein warmes Wasserbad können zur Unterstützung des Lösungsvorgangs verwendet werden.
Wirkungsmechanismus (Mechanism of Action)
| Targets/IC50/Ki |
EED
82 nM(Kd)
PRC2
114 nM(Kd)
|
|---|---|
| In vitro |
EED226 induces a conformational change upon binding EED, leading to loss of PRC2 activity. This compound also effectively inhibits PRC2 containing a mutant EZH2 protein resistant to SAM-competitive inhibitors. It regulates histone H3K27 methylation and PRC2 target gene expression in cells. In the in vitro enzymatic assays, this chemical inhibits PRC2 with an IC50 (half-maximal inhibitory concentration) of 23.4 nM when the H3K27me0 peptide is used as substrate and an IC50 of 53.5 nM when the mononucleosome is used as the substrate, with the stimulatory H3K27me3 added at 1 × Kact (1.0 μM). It is noncompetitive with both SAM and peptide substrate. This compound bound to EED and PRC2 complex with a 1:1 stoichiometry and Kd of 82 nM and 114 nM, respectively. It does not disrupt the PRC2 complex and could still occupy its binding pocket with a SAM-competitive EZH2 inhibitor bound to PRC2. This chemical shows remarkable selectivity for PRC2 complex over 21 other protein methyltransferases, kinases and other protein classes, The only other histone methyltransferase that can be inhibited by it is the EZH1-PRC2 complex. It with moderate permeability leads to a dose-dependent decrease of both global H3K27me3 and H3K27me2 markers in G401 cell.
|
| In vivo |
This compound effectively induces tumor regression in a mouse xenograft model. It in a solid dispersion formulation are well tolerated in animals. This chemical clearly demonstrates a dose-dependent efficacy in the mouse xenograph study. It inhibits the growth of diffuse large B-cell lymphoma (DLBCL) xenografts and reduces H3K27me3 levels to a similar extent as an EZH2 inhibitor. It has very low in vivo and in vitro clearance and approximately 100% oral bioavailability, low volume of distribution (0.8 L/kg), reasonable terminal t1/2 (2.2 h), and moderate plasma protein binding (PPB)(14.4%). Its solubility is relatively low and with little dependency on the pH of the medium.
|
Referenzen |
|

Die Produkte sind nur für Forschungszwecke bestimmt. Nicht zur Anwendung am Menschen. Kein Verkauf an Patienten.
©Copyright 2013 Selleck Chemicals. Alle Rechte vorbehalten.